Tampilkan postingan dengan label kimia. Tampilkan semua postingan

Selasa, 29 November 2011

Memasang VMD dari File Binary

Salah satu aplikasi kimia yang sering dipakai sebagai viewer dalam komputasi adalah software VMD.

"VMD is a molecular visualization program for displaying, animating, and analyzing large biomolecular systems using 3-D graphics and built-in scripting"

Untuk memasang VMD dalam Blankon/Ubuntu/Linux perlu dilakukan langkah-langkah berikut:

1. Download source code VMD dari website-nya yaitu http://www.ks.uiuc.edu/Research/vmd/vmd-1.9/files/final/vmd-1.9.bin.LINUX.opengl.tar.gz
2. Masuk ke terminal dengan klik Aplikasi --> Aksesoris --> Terminal atau tekan "Ctrl+Alt+T"
3. Masuk ke direktori tempat download file VMD
4. Ekstrak file tersebut dengan ketik: tar -xvf vmd-1.9.bin.LINUX.opengl.tar.gz
5. Masuk ke folder hasil ekstrak file VMD, ketik cd vmd-1.9
6. Baca file README dengan ketik: gedit README. Perhatikan bagian akhir file: Quick Installation Instructions
7. Perhatikan langkah-langkah instalasi berikut:

Quick Installation Instructions
-------------------------------------------------
Detailed instructions for compiling VMD from source code
can be found in the programmer's guide.

The Windows version of VMD is distributed as a self-extracting
archive, and should be entirely self explanatory.

The native MacOS X version of VMD is packaged as a disk image and is
extracted by opening the disk image, and dragging the "VMD" application
contained inside into an appropriate directory.

For quick installation of the binary distribution for Unix do the following:
1) Uncompress and untar the distribution into a working directory.
In this working directory, there are several subdirectories such
as bin, src, doc, data, as well as this README and a configure script.
Change to this working directory after the unpacking is complete.

2) Edit the file 'configure'; change the values for
the $install_library_dir and $install_bin_dir to a directory in
which vmd data files and executables should be installed, be sure
that you installing into a clean target directory and not overwriting
an existing version of VMD (which would otherwise give problems):

$install_bin_dir is the location of the startup script 'vmd'.
It should be located in the path of users interested in running VMD.

$install_library_dir is the location of all other VMD files.
This included the binary and helper scripts. It should not be
in the path.

3) A Makefile must be generated based on these configuration variables
by running "./configure".

4) After configuration is complete, cd to the src directory,
and type "make install". This will install VMD in the two
directories listed above. Note that running "make install"
twice will print error messages because you are attempting to
overwrite some read-only files. Similarly, if you have incorrectly
specified the target installation directories or attempt to overwrite
an existing VMD installation, you will get error messages.

5) When installed, type 'vmd' to start (make sure the
$install_bin_dir directory is in your path).

8. Baca file "configure", perhatikan bagian awalnya:

#!/usr/bin/env perl
# (C) Copyright 1995-2011 The Board of Trustees of the University of Illinois
# All Rights Reserved
#
# Usage: configure [ optionlist ] (See full option description down below)
# Perl 5.x must be in your path, and /usr/bin/env must exist in order
# for this to work correctly.

#################################################
# User modifiable installation parameters, can be overridden by env variables
#################################################
# Name of shell script used to start program; this is the name used by users
$install_name = "vmd";

# Directory where VMD startup script is installed, should be in users' paths.
$install_bin_dir="/usr/local/bin";

# Directory where VMD files and executables are installed
$install_library_dir="/usr/local/lib/$install_name";


# optionally override hard-coded defaults above with environment variables
if ($ENV{VMDINSTALLNAME}) {
$install_name = $ENV{VMDINSTALLNAME}
}
if ($ENV{VMDINSTALLBINDIR}) {
$install_bin_dir = $ENV{VMDINSTALLBINDIR}
}
if ($ENV{VMDINSTALLLIBRARYDIR}) {
$install_library_dir = $ENV{VMDINSTALLLIBRARYDIR}
}

9. Okay, sekarang masuk ke folder vmd-1.9
10.ketik: ./configure
11. masuk ke folder "src" dengan ketik: cd src
12. ketik: make install
13. Biarkan sampai instalasi selesai
14. untuk menjalankan software VMD ketik: vmd [Enter]
15. Selamat berlatih!

Selasa, 11 Oktober 2011

SOftware Kimia Avogadro

Avogadro


Avogadro is a beautiful molecule and material visualization and editing tool. It’s cross-platform and open source, produces high quality images and has some excellentscreencasts to help you get started.
Avogadro is an advanced molecule editor and visualizer designed for cross-platform use in computational chemistry, molecular modeling, bioinformatics, materials science, and related areas. It offers flexible high quality rendering and a powerful plugin architecture.
  • Cross-Platform: Molecular builder/editor for Windows, Linux, and Mac OS X.
  • Free, Open Source: Easy to install and all source code is available under the GNU GPL.
  • International: Translations into Chinese, French, German, Italian, Russian, Spanish, and others, with more languages to come.
  • Intuitive: Built to work easily for students and advanced researchers both.
  • Fast: Supports multi-threaded rendering and computation.
  • Extensible: Plugin architecture for developers, including rendering, interactive tools, commands, and Python scripts.
  • Flexible: Features include Open Babel import of chemical files, input generation for multiple computational chemistry packages, crystallography, and biomolecules.

Avogadro is an advanced molecular editor designed for cross-platform use in computational chemistry, molecular modeling, bioinformatics, materials science and related areas. It offers a flexible rendering framework and a powerful plugin architecture.
Avogadro is an application purely for scientists and students that have to draw three-dimensional molecular compositions that showing a molecule from every conceivable angle and perspective.
It works across platforms and is used in computational chemistry, molecular modeling, bio informatics, material science and other related areas. The aim has been to make 3D molecular manipulation as easy as possible and even if your not a scientists, it's quite entertaining just creating your own different coloured structures that look good. This can be done simply by using the mouse buttons and dragging the molecules on their three axis. Probably the best aspect of this program for those with some coding knowledge is that it can be easily extended and developed via plugins. Avogadro is based on Open Babel according to the developers and in future, will also offer scripting options. The main problem you'll find is that like with many open source collaborations, it is prone to bugs and errors so it may crash at any time.
Carbon Nanotube Rendered using Avogadro
Carbon Nanotube rendered using Avogadro

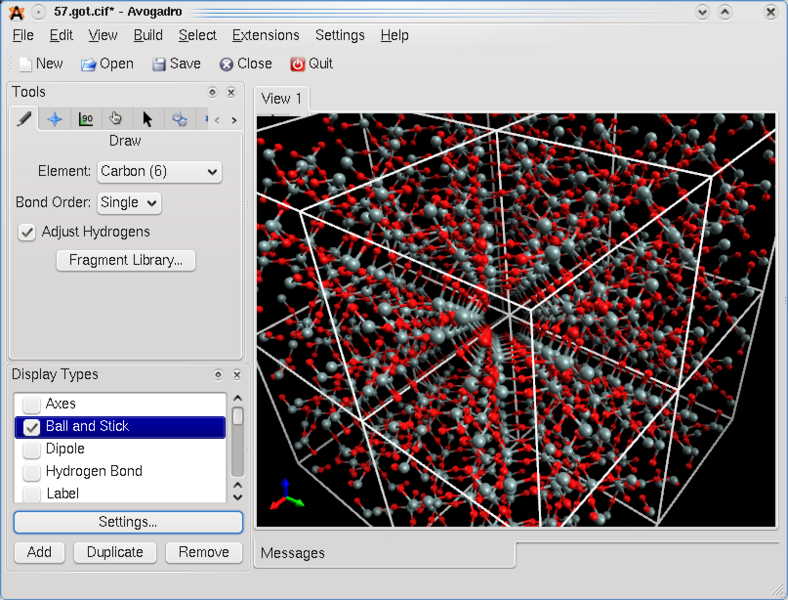
Unit Cell using Avogadro
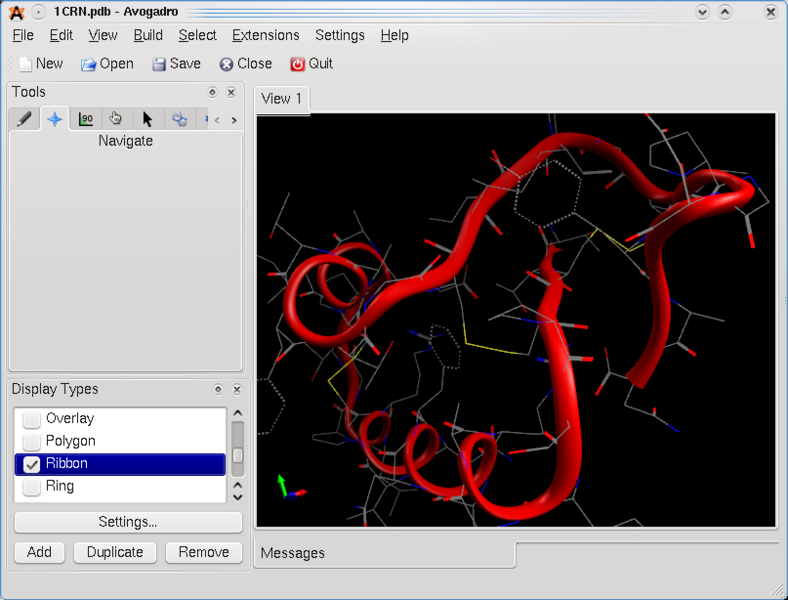
Protein Ribbon viewed in Avogadro

Molecular Orbital calculated using Avogadro
 

Sabtu, 01 Oktober 2011

Daftar Chemistry Software dari Sourceforge.net


Avogadro  Updated 2011-04-25

Avogadro is an advanced molecular editor designed for cross-platform use in computational chemistry, molecular modeling, bioinformatics, materials science and related areas. It offers a flexible rendering framework and a powerful plugin architecture.

Open Babel  Updated 2010-12-01

Open Babel is a chemical toolbox designed to speak the many languages of chemical data. It's an open, collaborative project allowing anyone to search, convert, analyze, or store data from molecular modeling, chemistry, biochemistry, or related areas

XDrawChem  Updated 2005-11-30

XDrawChem is a drawing application designed for drawing and analyzing chemical structures and reactions.

Gabedit  Updated 2011-09-29

Gabedit is a Graphical User Interface for Gamess-US, Gaussian, Molcas, Molpro, MPQC, OpenMopac, Orca, PC Gamess and Q-Chem computational chemistry packages.

SASHIMI  Updated 2011-09-21

The Sashimi project hosts the Trans-Proteomic Pipeline (TPP), a mature suite of tools for mass-spec (MS, MS/MS) based proteomics: statistical validation, quantitation, visualization, and converters from raw MS data to our open mzXML format.

Toxic Hazard Estimation  Updated 2011-08-05

A GUI application which estimates toxic hazard of chemical compounds. Provides a plugin framework to incorporate different approaches to the estimation. Platform independent (written in Java), with the use of The Chemistry Development Kit.

Bika Open Source LIS/LIMS  Updated 2011-07-25

Bika combines web content management and workflow processing for a one-stop web-based LIMS (Laboratory Information Management System) · Built in Plone · Professionally supported · Making waves · bi:ka (Zulu): tell it as it is

Zodiac  Updated 2009-09-03

Zodiac is a Molecular Modelling suite for drug design. Its features include support for haptic devices and nintendo wiimote fully undoable command list, MMFF94 implementation, POVRAY output.

QuteMol  Updated 2007-06-20

QuteMol is a real time, high quality molecular visualizer that offers an array of innovative visual effects. QuteMol aims at improving the clarity of rendered images and at offering a better understanding of the molecular 3D shape and spatial structure.

JSpecView Project  Updated 2011-09-20

The JSpecView Project provides JAVA applets for the display of JCAMP-DX and AnIML/CML spectral files. The source and binary files are available via the Code (sourceforge SVN) and Files menu options. See the WIKI pages (Hosted Apps) for more detail

DISCUS simulation package  Updated 2011-08-26

DISCUS is a set of programs to simulate the atomic structure of disordered materials and calculate experimental properties such as diffuse scattering or the atomic pair distribution function.

Chemitorium  Updated 2010-11-08

Free Organic Chemistry Analysis and Visualisation Tool; Chemical formula editor, calculation of threedimensional molecular structures, high-quality realtime rendering, ...

SASfit  Updated 2011-09-29

A program for fitting structural models to small angle scattering data.

GSim - tool for NMR spectroscopy  Updated 2011-05-26

GSim - free software tool for visualisation and processing of experimental and simulated nuclear magnetic resonance (NMR) spectra.

osra  Updated 2011-04-06

OSRA is a utility designed to convert graphical representations of chemical structures, as they appear in journal articles, patent documents, textbooks, trade magazines etc., into SMILES or SD file format- a computer recognizable molecular structure

Muse  Updated 2011-06-12

The aim of our project is to develop a simulation system to calculate the measurement uncertainty corresponding to the 'Guide To The Expression Of Uncertainty In Measurement' (GUM).

mzmatch  Updated 2011-09-21

mzMatch is a Java collection of small commandline tools specific for metabolomics MS data analysis. The tools are built on top of the PeakML core library, providing mass spectrometry specific functionality and access to the PeakML file format.

OpenThermo  Updated 2009-08-15

OpenThermo is a program package for statistical thermodynamics computations beyond rigid rotor - harmonic oscillator approximation

cutenmr  Updated 2010-11-09

Free, multi platform NMR processing application that reads Bruker and Varian files.

bni-tools  Updated 2010-09-29

The BNI (Beyond Normal Interaction)- Tools is a plug in for the PyMOL molecular visualization system which adds additional functionalities and presets to the PyMOL GUI and also adds useful extended commands.

MESMER  Updated 2011-02-18

MESMER (Master Equation Solver for Multi Energy-well Reactions) models the interaction between collisional energy transfer and chemical reaction for dissociation, isomerisation, association, and non-adiabatic hopping processes.

Impedance Analysis and Instrumentation  Updated 2011-03-16

Electrical Impedance Tomography and Impedance Spectroscopy software that I am developing for my PhD project.

ED Software  Updated 2009-08-19

ED Software project contains several programs used (mostly) for processing gas-phase electron diffraction (GED) experimental data.

Screening Assistant 2  Updated 2011-07-25

ScreeningAssistant 2 is a modular software dedicated to perform various simple and advanced chemoinformatics analysis around chemical libraries.

zephyr  Updated 2009-09-23

A script-driven measurement automation program for applications involving various measurement instruments and actuators. Supports instruments from Agilent, Eco Physics, ICS Electronics, Keithley, Omega, Teledyne and other manufacturers.

pteros  Updated 2011-06-09

Pteros is the C++ library for custom molecular modeling and simulations codes designed for researchers, not for C++ gurus. Provides facilities for PDB, XTC and TRR files IO, powerful selections, geometry transformations, RMSD fitting and alignment, etc.

sdf2xyz2sdf  Updated 2011-03-12

An open-source software to convert SDF files into TINKER XYZ files (and viceversa) with automatic assignment of MMFF94 atom types, bond types and charges

PLASMAKIN: a chemical kinetics package  Updated 2009-03-31

PLASMAKIN is a package to handle physical and chemical data used in plasma physics modeling and to compute gas-phase and gas-surface kinetics data: particle production and loss rates, photon emission spectra and energy exchange rates.

EvoGrid  Updated 2010-08-06

The EvoGrid is an effort to develop a distributed simulation system to simulate potential pre-biotic chemical environments.

greylag  Updated 2009-11-02

Tandem mass spectral peptide identification and validation software, similar to X!Tandem, OMSSA, MyriMatch. Suitable for single hosts through large clusters. Written in Python for simplicity, with performance-critical sections in C++.

NOSE  Updated 2009-02-02

NOSE is a package for simulation of linear and non-linear optical and infrared spectra, including absorption, fluorescence, circular dichroism, pump-probe, photon echo and other.

Quantum Simulation Software  Updated 2005-04-06

qsims simulates quantum dynamics with an (almost) arbitrary time-dependent Hamiltonian. qsims uses a Chebychev polynomial expansion of the Schrodinger propagator, and the pseudospectral method developed by R. Kosloff.

Cantharella  Updated 2011-08-19

The information system (IS) “Cantharella: Pharmacochemical database of natural substances” share and sustain pharmacochemical data of all organisms collected for the study of their natural substances, with a controlled access via internet.

FlameBitmaps  Updated 2009-06-10

Calculating 2D-Gray-scale bitmaps Calculation of a 3D-rotation-body form a detected area by using different image processing algorithms. Developed for the German Aerospace Center (DLR). Used Projects: Substance (GUI)

facile  Updated 2011-08-28

A command-line biochemical network compiler for systems biology.

GlycanEditor

A visual editor for the definition of Carbohydrate structures, computation of masses and derivation of fragments.

PetitChimiste

PetitChimiste is a 3D presentation tool for molecules. Features: - Load Molec3D,Alchemy,PDB files. - Calculations (angles and lengths) between atoms. - Wiews: Wireframe & Solid in true colors. - Portable on all platforms

SonicData  Updated 2008-01-23

SonicData is a data collection and analysis program used in connection with a sound card voltage-to-frequency interface. Its intended use is for laboratory instrumentation.

SpOK - Spectrum Organisation Kit  Updated 2005-11-04

SpOK - The Spectrum Organisation Kit, a program for the organisation and visualisation of spectral data written in Java. Sorry - this project is not being continued. All functionality was integrated into the Bioclipse project...

QMcBeaver  Updated 2008-04-29

QMcBeaver is an object-oriented program to perform Quantum Monte Carlo calculations on atoms and molecules. It is designed to be easy to modify, allowing new ideas to be quickly implemented.

SPAS  Updated 2005-12-18

SPAS (Structure Property Analysis Software) is a Quantitative Structure-Property Relationship (QSPR) software.

GAST-Viz - Atomistic Simulation Toolkit  Updated 2004-03-11

GAST-Viz is a toolkit allowing for the creation of OpenGL atomistic simulation applications using discrete element modeling methods. A number of sample applications are included.

Vimes  Updated 2009-08-26

Vimes: Visual interface to materials simulations - An open source, python/Open GL interface to molecular and materials simulation programs

Chamber Interior Ballistics  Updated 2006-04-20

chamber is a reactive hydrodynamic modeling tool specifically designed for small arms interior ballistics research. Multiple equations of state and burn models are supported. chamber is an adaptation of the BKW, SIN, 2DL and 2DE codes in one package.

runsbml  Updated 2005-06-07

runsbml - Pathway Simulation Tool emulates the dynamics of biological pathways by calculating concentrations of biochemical compounds at different times and conditions.

Open ESR

A set of Fortran 77/95 libraries and drivers for the analysis of CW/Pulsed Electron Paramagnetic (Spin) Resonance spectra.

Mops/Sweep Particle Reactor Solver  Updated 2006-05-18

A reactor solver which uses stochastic particle methods to model particle population balances. This code is developed by the CoMo group in the chemical engineering department at the university of Cambridge (como.cheng.cam.ac.uk).

TinyHF

A lightweight implementation of molecular Hartree-Fock theory with automatically generated integration routines, extended Hueckel initial guess, DIIS convergence acceleration, modified Mulliken population analysis, etc.

spectratools

An open-source library to retrieve data from infrared (IR) spectroscopy data files. Having the data in a plain-text format make it simple to use in other applications; for example, making a plot ready to be submitted as part of a journal article.

ARVE

The Atmosphere-Regolith-Vegetation (ARVE) project has the goal of creating, maintaining and distributing a state-of-the-art Dynamic Global Vegetation Model (ARVE-DGVM) for investigating the role of the Earth's land surface in the climate system.

CheMecher

CheMecher is a fully-featured chemical simulator that supports an arbitrary number of reaction steps and chemical species. Uses the Runge-Kutta algorithm for numeric solutions.

Force Field X

FFX is a molecular biophysics tool written in pure Java. We have implemented the Atomic Multipole Optimized Energetics for Biomolecular Applications (AMOEBA) force field with special features for crystallography.

ABFF

Project for develop force fields with using ab-initio calculations

GEMS

Grid-Enabled Molecular Simulation Repository (GEMS)

Instans  Updated 2010-12-01

Methods and testing of methods for automatic analysis of in situ cyclic voltammetry data. This, at least initially, is the code from my masters thesis, which was done as a contribution to a larger project called Aevum. Aevum is being developed at t

SDFCherry

SDFCherry is a command-line program that searches for textual matches in molecular structure files of SDF format and outputs the structures that match. Optionally adds a data field to the matched structures, e.g., biological activity. Useful for QSAR.

cca-forum  Updated 2011-05-07

Cca-forum unifies the Common Component Architecture tools and tutorial. It includes the CCA specifications, the Ccaffeine framework for HPC, and related tools. These support multilanguage scientific and parallel computing.

Kitchen Sink

Run-time libraries to support the Kitchen Sink, an all-purpose chemical process control board, based on the Atmel AVR ATmega1280 chip, using the Arduino core libraries.

XStar  Updated 2010-06-30

A Collection of STAR/CIF processing libraries and related tools, including sub-projects: AStar, TEStar and others...

bruread

bruread is a program to read and display NMR data in the Bruker XWIN-NMR format. The program has a command-line interface and a graphic display based on gnuplot. - Note that this is only an NMR spectrum *viewer*, not a full-fledged data evaluation suite

hr

hr ("High Resolution") is a small command-line utility to calculate possible elemental compositions for a given mass. It allows to run calculations interactively, in batch mode, via the command line, or (using a webserver) via a web interface.

isotope

'isotope' is a small command-line utility to calculate the isotope pattern for a given chemical formula. It allows to run calculations interactively, in batch mode, via the command line, or (using a webserver) via a web interface.

cdfread

cdfread is a program for people wirking with mass spectrometry datasets. cdfread implements the routines to read mass spectra and mass chromatograms from data files in netCDF ("Andi-MS") format. Centroid and profile data are supported.

The Chemicals Database

The Chemicals Database is a simple yet effective approach to track a stock of products. It is accessible from any workplace over Intra/Internet (http). It was designed in particular to track chemicals in a laboratory, but is suitable for other items.

The Analyses Database

The Analyses Database is a collection of software to archive and to track rawdata files from analytical instruments. The system is set up using freely accessible tools - MySQL, apache, PHP, Linux. Several utilities for MS Windows are provided.

remap

A matlab/octave based reaction transport model describing microbially mediated isotope fractionation processes in porous media.

CARMEL 'CARbon MEchanics Lab'

CARMEL is a suit of parallel codes designed to perform classical molecular dynamics simulations of atomic and ionic systems Potentials : Lennard Jones, Yukawa, Tersoff Bond order Integrators : NPT, NVT, NVE

3DMolvis  Updated 2009-08-03

3DMolvis is a tool designed to make easy learning of basic structure and function of biological molecules.

dmcut2

dmcut2 is a scientific program designed to simulate chemical reactions at surfaces using Metropolis and Monte-Carlo methods. Short ranged interactions between adsorbates may be taken into account. Emphasis was put on code readability and usability.

platestats

The platestats software aims to provide a convenient solution for analyzing the results of biological assays performed in 96- and 384-well plates.

Adaptive Biasing Force

This is an implementation of the adaptive biasing force method (ABF) in NAMD. The code for ABF was developed by Eric Darve, Stanford University. ABF is a method created by E. Darve and A. Pohorille to calculate the free energy of bio-molecules.

Olex2

Olex2 is visualisation software for small-molecule crystallography developed at Durham University/EPSRC. It provides comprehensive tools for crystallographic model manipulation for the end user and an extensible development framework for programmers.

Algorithms in Spectroscopy  Updated 2008-06-14

The aim of the project is to provide open source collection of algorithms in the field of spectroscopy: data handling and processing, modeling and artificial intelligence tools.

wyckconf

wyckconf is a tool to generate crystal structure configurations from Wyckoff coordinate input, and to regenerate symmetries in the resulting structures of molecular dynamics calculations.

spcview  Updated 2008-03-08

The software "SPCView" is used for XRF qualitative analysis of various substances with "SPECTROSCAN MAKC" spectrometers.

Strigi-chemical

Bio- chemical meta data extraction, indexing and search for desktop search engine Strigi

MitoMAS_membrane  Updated 2007-12-04

Mito-MAS-m is a simulator of the mitochondrial inner membrane and the enzymatic complexes embedded in it, implementing a coarse-grained (CG) model of the molecules using rigid structures and Dissipative Particules Dynamics (DPD) as motion equation.

Tools for SAXS Simulation

A family of programs for building coarse-grained models (eg. of macromolecules), manipulating them (geometrical transformations and more) and calculating their X-ray scattering patterns (1D and 2D too). Uses PVM for parallelization.

CT8K

CT8K is an experimental quantum chemistry program intended to bring state of the art methods to the desktop PC

ChemRuby

ChemRuby is a framework for developing cheminformatics applications in Ruby. It will let you retrieve chemical information from variety of data sources, fast substructure search, draw a chemical structure, and calculate chemical properties.

molmass

molmass is a cross-platform python program usable from the gui or commandline, for calculating the molar mass of a chemical formula.

HoloJ

A suite for Transmission Electron Microscopy holograms reconstruction and analysis

Quantitative Chirality

Quantitative Chirality: A multiplatform implementation of the 1995 Osipov, Pickup and Dunmur algorithm for calculating the chirality index and predicting helical twisting power.

Tools for ab initio calculations  Updated 2006-04-25

Soft for ab initio and MD simulating of water system. Contains program for ICE: construct ice, specific monte-carlo simulation, proton rearrangement calculations; for PDB-water: calculate rotation energy profile, for Effective Fragment simulations.

Openbabel Extensions  Updated 2006-01-30

OBExtensions is a extension library for the chemoinformatics library Openbabel. It provides extensions to Openbabel, being mostly GUI widgets. OBExtensions has a 2D/3D viewer, 2D coordinates generator and some other extensions.

The Blue Obelisk

The Blue Obelisk is a group of scientists driven by a belief in Open Source, Open Standards and Open Data, expressed in code, data, algorithms, specifications, tutorials, demonstrations, articles and anything that helps get the message across.

Olive  Updated 2011-09-14

Sistema de informacion Nutricional - Olive. Desarrollado por LandSoft S.A.
FORM ISIAN DATA MAHASISWA Untuk pendataan mahasiswa peserta mata kuliah "Program Komputer untuk Pengajaran Kimia Berbasis ICT", silahkan anda ikuti/klik link berikut:

FORM ISIAN DATA MAHASISWA

Terima kasih atas partisipasi anda! Semoga sukses selalu untuk anda!